Find out where you actually stand
Most quality managers I meet already suspect something isn't right. What they can't do is put a number on it, or say which of the twenty things on their list is the one that will hurt them.
That's what an assessment is for. I go through your quality system against the regulations that apply to you, and I tell you where the gaps are and which ones matter. You get a document you can take to your board, or your parent company, or the person holding the budget.
It's a fixed piece of work with a clear end. Plenty of clients take the report and act on it themselves.
I work with pharmaceutical manufacturers, sponsors and clinical research organisations, in Australia and overseas.

28 years in life sciences across three continents
Experience with USFDA, TGA and MHRA audits
What you get out of it
A ranked list, not a long one.
I've seen assessments come back with sixty observations and no indication of which three are the most critical to act upon. That isn't useful to anyone.
You'll know which gaps are regulatory and which are just costing you time and money.
Those get treated differently, and most reports don't separate them.
You'll usually find things you can stop doing.
That's often where the quickest return is, and it almost never appears on a gap analysis, because gap analyses are built to find what's missing.
And you'll be able to answer the question "where do we stand" without a fortnight of preparation.
That matters more than it sounds. It's most of what inspection readiness actually is.
Why the same problems keep coming back
If your findings repeat, it's almost never because people aren't trying.
I was doing a facility assessment recently and found four areas where operators weren't replacing a dirty mat outside the room. When I looked into it, every mat change had to be recorded in a log. During processing the operators were moving in and out constantly, and the logging just added time without adding value. So the mats stayed down, and became a source of contamination rather than a control against it.
No regulation requires mat changes to be logged. Nobody on site knew why the log existed or when it had been introduced. Compliance was being held back by over-documentation.
That's the pattern. The system gets built to survive an audit rather than to run a business. It looks correct on paper, and in practice people find a faster way. Then each finding gets its own fix, and someone adds a form, or a second signature, or another procedure. Every one of those decisions was sensible at the time. Nobody has ever gone back and looked at what all of them weigh together.
Meanwhile the procedures were often written by people who don't do the task, so they describe a version of the work that doesn't exist.
This is the pattern I keep running into, and it's why I work the way I do.
You'll recognise some of these
The same findings turn up every audit cycle
Two or three people are holding compliance together
Preparation starts when the inspection date lands
Your staff have their own workarounds and you half know about them
CAPAs close on time and nothing changes
Nobody could tell you today where you stand
If more than a couple of those are familiar, it's systemic. Trying harder won't fix it.
What I look at
Your documented system first: procedures, records, governance, and what your regulatory position actually demands of them.
Then the system as it runs. I map the processes, walk the facility, and interview the people doing the work rather than only speaking to QA. I'm looking for what they understand, what they're actually doing, and what's getting in their way. That part is the assessment. Everything else is reading.
The gap between those two is where most findings come from, and a desk review can't see it.
I'll also tell you where you're over-controlled, because too much control is a real problem and almost nobody goes looking for it. Extra approvals slow decisions. Extra forms get completed late, or after the fact. A procedure nobody can follow gets ignored. All three eventually turn up as findings.
Every finding comes with the reason behind it
This is the part that makes the difference between a report and something people act on.
For every gap I identify, I explain why it would raise a red flag for an inspector. Not just what's missing, but what the regulator is actually worried about and what they'd ask if they were standing there.
That matters because the most common reaction to a finding isn't resistance, it's bewilderment. People have been working a certain way for years and can't understand why it's suddenly a problem. Handing them a list of gaps doesn't fix that. Explaining what sits behind each one does, and it means the next decision they make without me is more likely to be the right one.
What you receive
A findings report.
For each area: the regulatory clause that applies, where you currently sit against it, and what's missing.
A criticality rating on every gap.
So you can work through them by risk, not by the order they happen to appear in the report.
Short-term and long-term actions, kept separate.
What needs dealing with now, and what belongs in next year's plan. If you mix them, the urgent things get lost.
A walkthrough with your team.
I take the people who have to act on the findings through them. Emailing a PDF and hoping isn't an outcome.
If you have an inspection coming, or findings to close out
Two different situations. Same work underneath.
Before. I look for the system issues most likely to be raised, and we deal with them while there's still time. Ranked by what an inspector will actually chase, which isn't always what's easiest to fix.
Most preparation starts too late because it's driven by the date rather than by the risk. And a lot of it is tidying rather than testing. Documents get put in order and nobody checks whether the person who has to explain them can.
After. If you've already been inspected and you've got findings to address, that's the same assessment work with a deadline attached. I'll help you get behind each finding, work out what has to change so it doesn't come back, and bring your team along rather than leaving them with a task list they don't understand.
I don't attend inspections. My work sits either side of the day itself.
Where this leads
An assessment tells you where you stand.
If what I find is structural, rebuilding or simplifying the system is a bigger piece of work, and it has its own page.
If you want to know how I approach any of it: understand the operation first, take out what isn't doing any work or isn't compliant, then implement and verify what's left against the requirements.
How you'd know it was worth doing
Whether the same findings come back next cycle.
How long it takes you to answer "where do we stand".
Whether closing a CAPA actually changes anything.
And whether people are using the documented process or their own version of it.
What those numbers become depends on what you do with the report.
Proof
After a TGA finding — multi-product medicines manufacturer
A TGA audit found major gaps in the rationale behind controls at a medicines manufacturer. Staff could not understand why long-standing arrangements were now non-compliant, and QA and operations disagreed on implementation. I reviewed the finding and documents, walked the facility, and interviewed staff. The resulting report prioritised each gap, explained why it would concern an inspector, and proposed mitigations. Tailored training helped both teams work from the same understanding.
eQMS effectiveness review — pharmaceutical CDMO
A pharmaceutical CDMO had implemented an eQMS quickly during the pandemic. Manual controls still ran alongside it, and the people who built it had left. I assessed the system against PIC/S data integrity guidance, mapped its processes, and delivered tailored QA training. The assessment exposed compliance and data integrity gaps before a regulator did. A short- and long-term plan helped the client prepare to replace the system and put remedial actions in place to hold the compliance position in the meantime.
Batch release, unblocked — contract development and manufacturing organisation
A CDMO’s final batch review and release had become a bottleneck. Recurring documentation errors, different QA and production priorities, and inconsistent understanding of compliance slowed the process. I assessed the process and brought both teams together to map it, design a batch review procedure, and train staff. The joint approach exposed duplicated steps and practical constraints. Redundant activities were removed, timelines set for each stage, and the project deliverable timeline was met.
What clients say.
“What stood out most was the ability to translate complex GMP requirements into clear, actionable recommendations that were meaningful for both Quality and Operations. The sessions were not just about identifying gaps, but also about building awareness, ownership, and long-term sustainability.”
Shalin Parikh, Head of Quality, Mayne Pharma
- 1. A conversation.
- 2. The assessment.
- 3. Analysis.
- 4. The report and walkthrough.
- 5. Help afterwards, if you want it.
How it runs
1. A conversation.
A short call so I understand the situation. You'll get an honest view on whether an assessment is the right starting point, and I'll tell you if it isn't. Thirty minutes and some candour about what isn't working.
How it runs
2. The assessment.
Document review, process mapping, a walk through the facility, and interviews with your people. I'll need access to procedures and records, and time with staff.
How it runs
3. Analysis.
I rank the gaps by regulatory risk and by what they're costing you operationally. Findings split into what's mandatory and what's a choice. Nothing needed from you here.
How it runs
4. The report and walkthrough.
Short-term and long-term actions, taken through with your team, with the reasoning behind each finding. You end up with a plan you could run without me.
How it runs
5. Help afterwards, if you want it.
Some clients take the report and run it themselves. Some ask me to stay on. Either is fine, and the assessment is finished either way.
How long the whole thing takes depends on your size and how many systems are in scope. I'll give you a rough estimate on the call and a firm one in the proposal.
Who you'd be dealing with
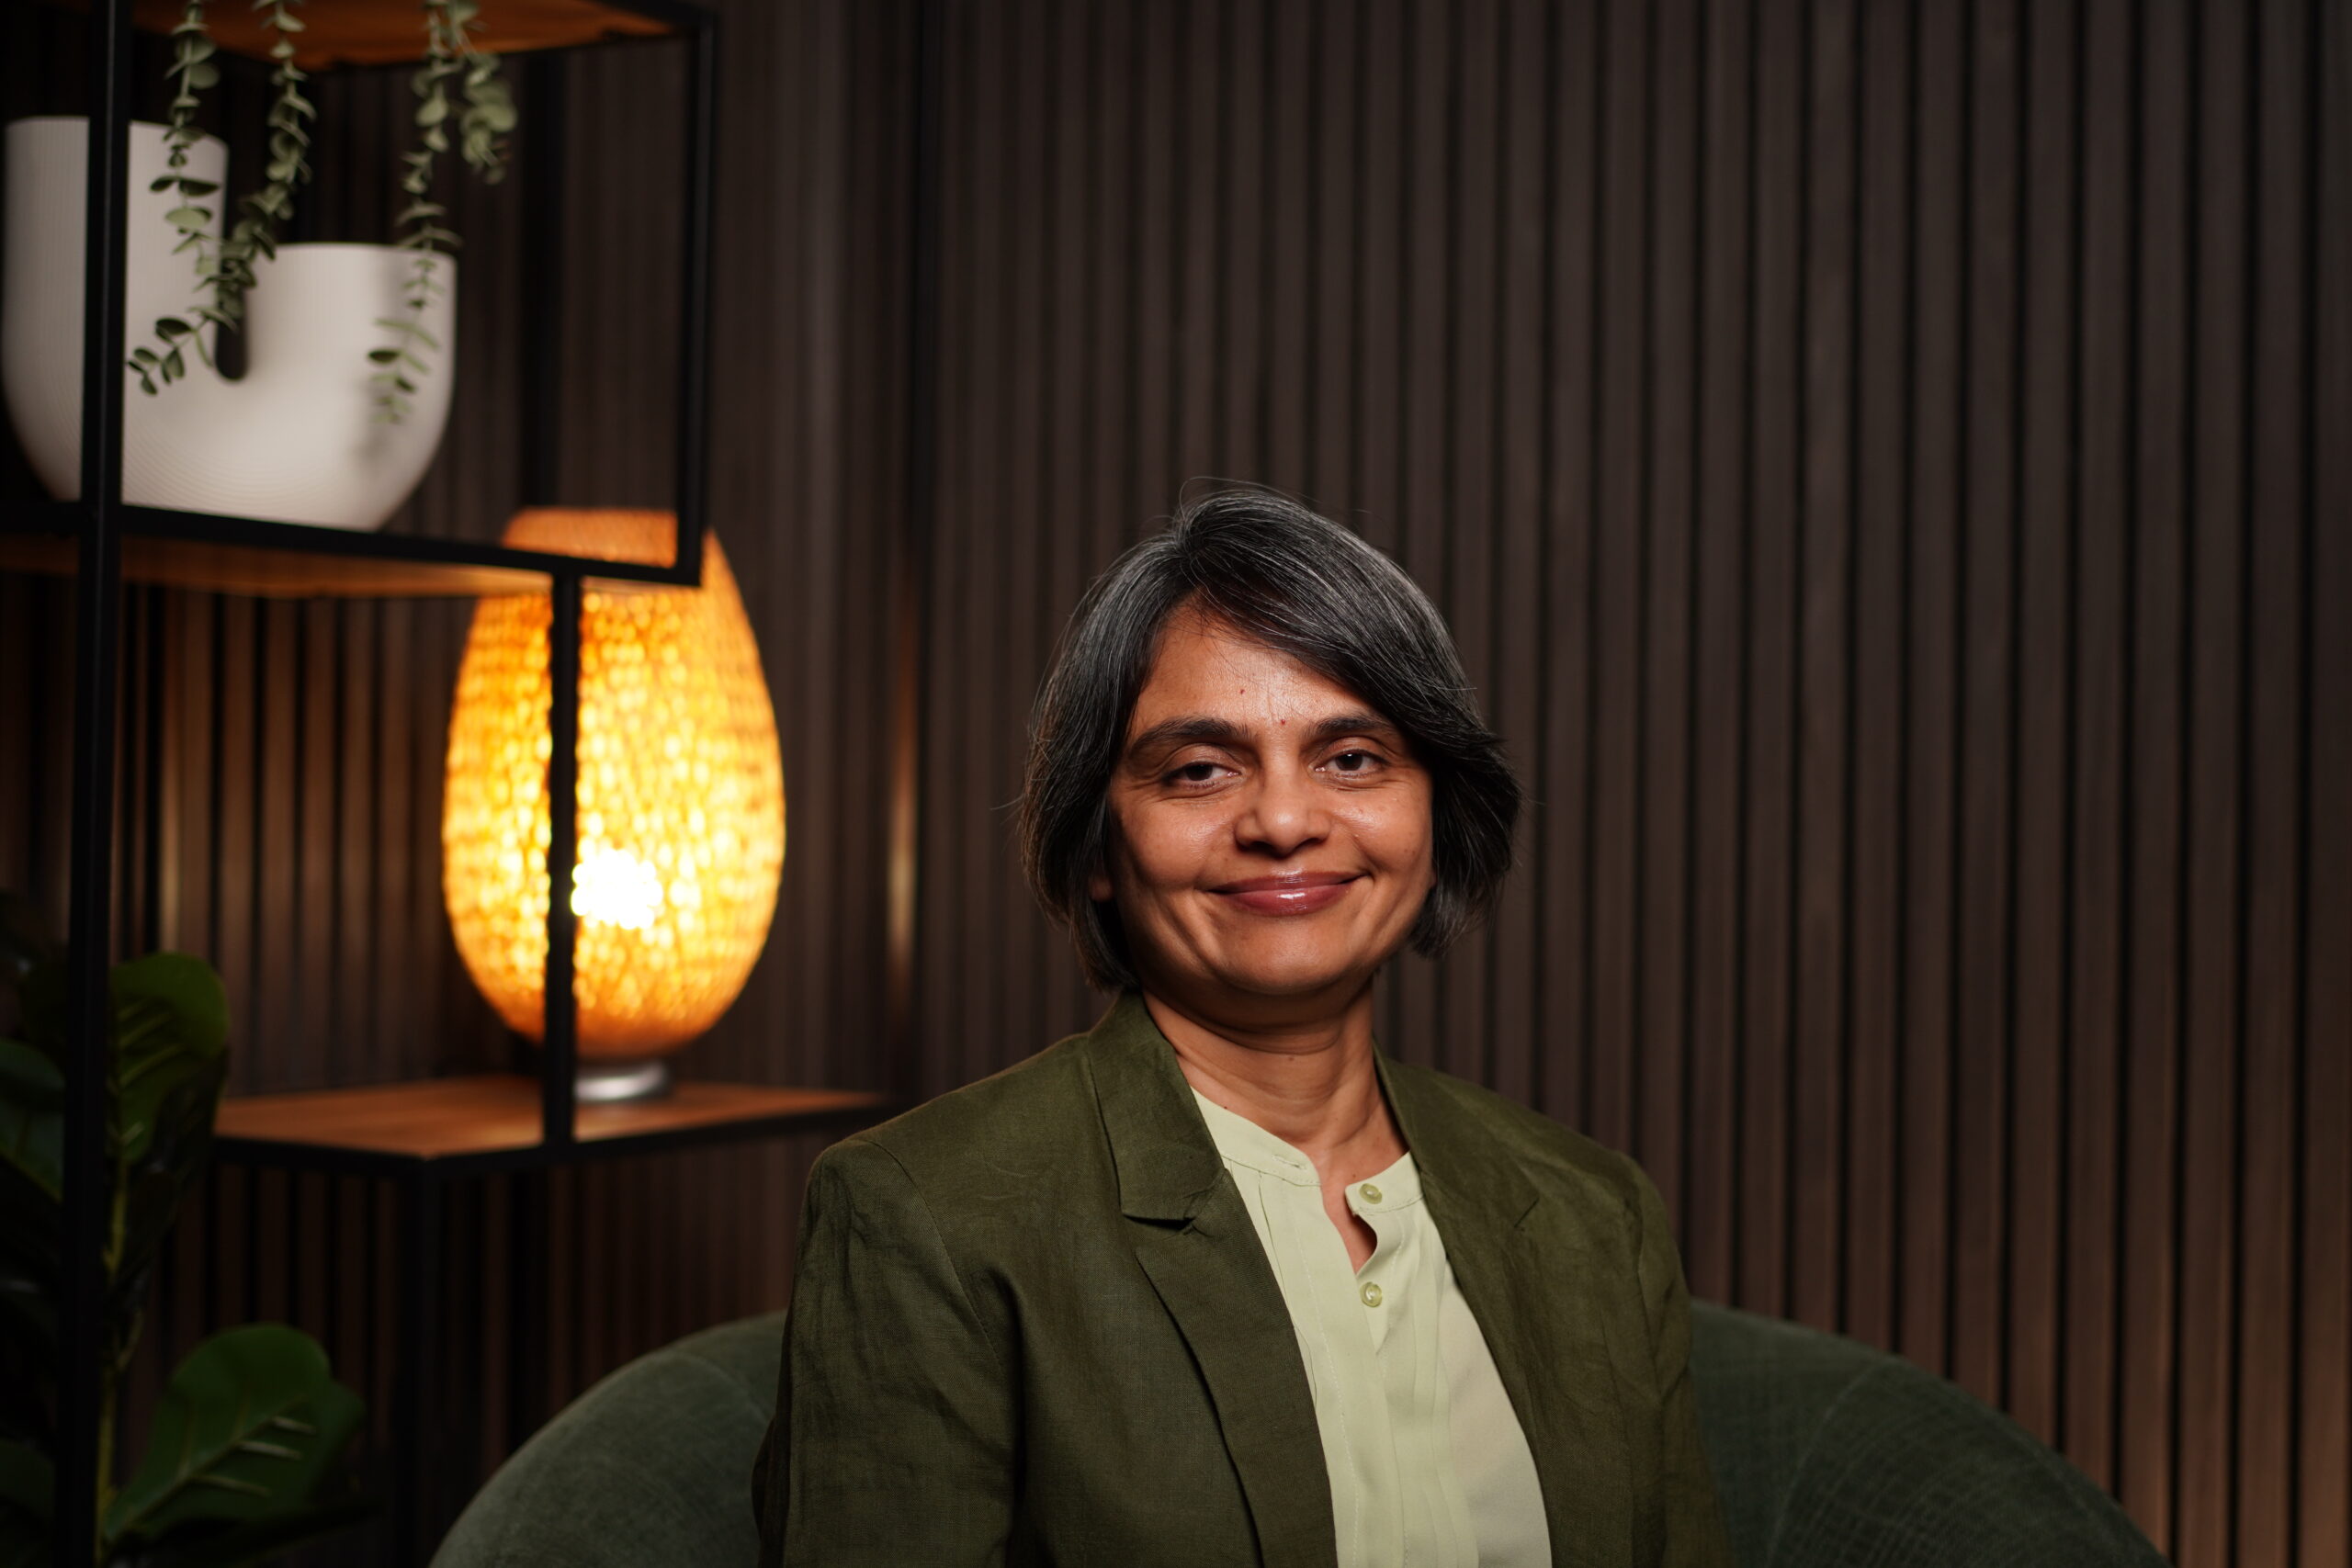
I'm Antiksha Joshi. I've spent 28 years in pharmaceutical and life sciences quality across three continents. I am Chair of Women in Pharma, ISPE Australasia Affiliate.
I also write and speak on quality systems for various knowledge sharing organisations and online publications.
I've worked in and around systems that have been through USFDA, MHRA and TGA scrutiny. My background is in microbiology. I also hold a diploma in counselling, which sounds like an odd combination until you've watched a technically perfect quality system fail because nobody could be persuaded to use it.
Years ago I was on a team investigating a major batch failure. Operations would suggest a fix, and I'd ask questions until I understood how the work actually got done on the floor, then explain why their suggestion wouldn't hold and what a regulator would ask if we went ahead with it. At the end of it, one of them said nobody had ever explained why they needed to do something before, and that now it made sense.
I've worked that way ever since.
Questions people ask before they call
What does the assessment actually cover?
Your quality system as written and as it's actually run, against the regulations that apply to you. Both halves. The second one is where the useful findings are.
How long does it take?
It depends on your size and how many systems are in scope. I'll give you a rough estimate on the call and a firm one in the proposal.
What do we get at the end?
A findings report with the applicable clause, where you sit against it, the gap, a criticality rating, and short and long-term actions. Plus a session walking your team through it.
Do you explain why something is a finding, or just that it is?
Always why. A list of gaps without the reasoning behind them doesn't change how anyone works, and it means every future decision has to come back to me.
How much of our people's time will it take?
Some, and I won't pretend otherwise. I need time with the people who do the work, not just with QA. That's what separates an assessment from a document review.
GxP is broader than GMP. Does that mean you cover us?
GxP covers clinical and laboratory practice as well as manufacturing. Most of my work is GMP and manufacturing. Tell me your situation on a call and I'll tell you honestly whether I'm the right person.
Do we have to act on what you find?
No. It's your report. Some clients work through it themselves, some ask me to help, and some put it aside until the timing is better.
Can you help us fix it?
If you want me to. It's a separate piece of work and a separate decision. The assessment is complete on its own.